Contents Menu Expand Light mode Dark mode Auto light/dark, in light mode Auto light/dark, in dark mode Skip to content
U-Chrom 0.3 (dev)
Light Logo Dark Logo
U-Chrom 0.3 (dev)

Getting started

  • Installation
  • Quick start
  • Tutorial gallery
  • Concepts
  • Datasets

By data type

  • Chromatin tracing
    • Reading tracing data
      • Importing FOF-CT chromatin-tracing data
      • Importing PyHiM chromatin-trace tables (ECSV)
    • From detections to traces
      • Spot-to-trace alignment with jie
    • Filling missing loci
      • Imputing missing FISH coordinates
    • Calling structures
      • Loop calling on chromatin tracing data
      • TAD calling
      • Single-allele domains with FISHnet
      • A/B compartments from chromatin tracing
  • Imaging multi-omics: DNA seqFISH+
    • Reading seqFISH+ multi-omics data
      • Importing seqFISH+ multi-omics data (Takei 2025 cerebellum)
    • Marks, cells and tissue
  • Single-cell and spatial Hi-C
    • Contact maps per cell
    • Pseudo-bulk maps
      • Pseudo-bulk Hi-C: compartments per cell type, at matched depth
    • 3-D structures of single cells
      • Single-cell 3-D genome reconstruction: NucDynamics and EMber
    • Cell embeddings
      • Contact-map cell embedding with FastHigashi
      • Cell embeddings and clusters across modalities
  • Bulk Hi-C
    • Calling structures from a map
      • Calling structures from a bulk Hi-C map
    • Structures from a bulk map: MDS and IGM
      • Bulk Hi-C to 3-D: MDS and IGM population deconvolution
    • Hi-C with imaging: GEM-FISH
      • Joint Hi-C + FISH reconstruction with GEM-FISH

Common to all data

  • Data model & storage
    • ChromData: the data model
    • The .chromdata.zarr store
    • ChromData
    • Linked Multi-Omics
  • Per-locus features
    • Per-locus features
  • Visualisation
    • Visualisation: plots and the web browser
    • The web browser
    • The data atlas

Reference

  • API reference
    • chromdata (the data model)
    • uchrom.tl / uchrom.pp
    • uchrom.datasets
    • uchrom.io
    • uchrom.im
    • uchrom.recon
    • uchrom.recon.fish
    • uchrom.strc.loop
    • uchrom.strc.tad
    • uchrom.strc.comp
    • uchrom.strc.enrichment
    • uchrom.fea
    • uchrom.emb
    • uchrom.pl
    • uchrom.utils
    • uchrom_browser (the web browser)
  • File formats
  • ChromData Specification
  • Datasets: sources and recipes
  • ChromData 2.0 Design
  • Development
  • Changelog
  • License
Back to top
View this page
Edit this page

Tutorial gallery

Every tutorial, by data type and analysis step — each runs on real data, from a dataset to its results. Click a card to open the tutorial; the chapters (tracing, seqFISH, single-cell Hi-C, bulk Hi-C) explain the methods around them.

Chromatin tracing
Importing FOF-CT chromatin-tracing data4DN FOF-CT tracing with its cell and RNA tables, read and written backImporting PyHiM chromatin-trace tables (ECSV)PyHiM ECSV traces (Bintu 2018 IMR90) as a ChromDataSpot-to-trace alignment with jieRaw seqFISH+ spots to chromosome traces, against the authors' tracesImputing missing FISH coordinatesFilling missing loci — linear, cubic, SnapFISH-IMPUTE — scored on hidden spotsTAD callingTADs from 3-D distances (ArcFISH), next to Hi-C domains of the same lociSingle-allele domains with FISHnetDomains of single alleles (FISHnet) and how often each boundary is usedLoop calling on chromatin tracing dataLoops from 3-D distances with the axis-wise F-testA/B compartments from chromatin tracingA/B compartments from imaging, against Hi-C of the same cells
Imaging multi-omics: DNA seqFISH+
Importing seqFISH+ multi-omics data (Takei 2025 cerebellum)DNA seqFISH+ with chromatin marks and RNA (Takei 2025 cerebellum)

Single-cell and spatial Hi-C

Pseudo-bulk Hi-C: compartments per cell type, at matched depthCell-type maps summed from an atlas store over HTTP; compartment strength at matched depthSingle-cell 3-D genome reconstruction: NucDynamics and EMberSingle-cell 3-D structures with NucDynamics and EMber on the native engineContact-map cell embedding with FastHigashiCells embedded by their contact maps (FastHigashi)Cell embeddings and clusters across modalitiesCells by RNA, ATAC or contacts (scHiCAR), and imaged cells by chromatin marks

Bulk Hi-C
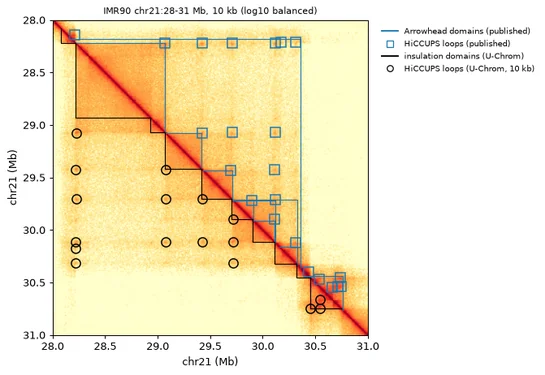
Calling structures from a bulk Hi-C mapCompartments, insulation domains, HiCCUPS / Mustache loops and APA, against the published callsBulk Hi-C to 3-D: MDS and IGM population deconvolutionA consensus structure (MDS) and a population of structures (IGM) from one mapJoint Hi-C + FISH reconstruction with GEM-FISHHi-C and FISH distances in one model (GEM-FISH)

Common to all data
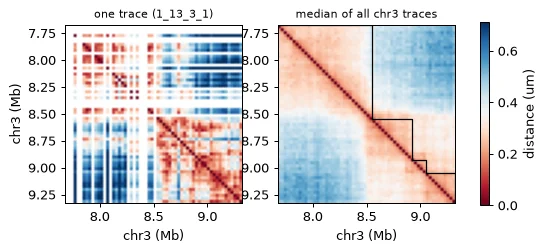
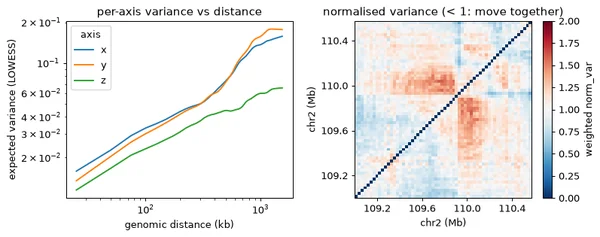
ChromData: the data modelChromData: bins, spots, tracks, intervals, results, subsettingThe .chromdata.zarr storeThe .chromdata.zarr store — backed and remote reads, streaming, linked and embedded dataPer-locus featuresDistance maps, variance features, peaks and projections along the genomeVisualisation: plots and the web browserPlots with uchrom.pl and the web browser from Python
Next
Concepts
Previous
Quick start
Copyright © 2026, Weize Xu
Made with Sphinx and @pradyunsg's Furo
On this page
  • Tutorial gallery
    • Chromatin tracing
    • Imaging multi-omics: DNA seqFISH+
    • Single-cell and spatial Hi-C
    • Bulk Hi-C
    • Common to all data